Manufacturers established outside the EU generally need to appoint an EU Authorised Representative before placing medical devices or in vitro diagnostic medical devices on the EU market. Under the MDR and IVDR, the representative has specific regulatory responsibilities and may be jointly and severally liable with the manufacturer for defective devices under the conditions set out in Article 11(5).
What is an EU Authorised Representative (EU-REP)?
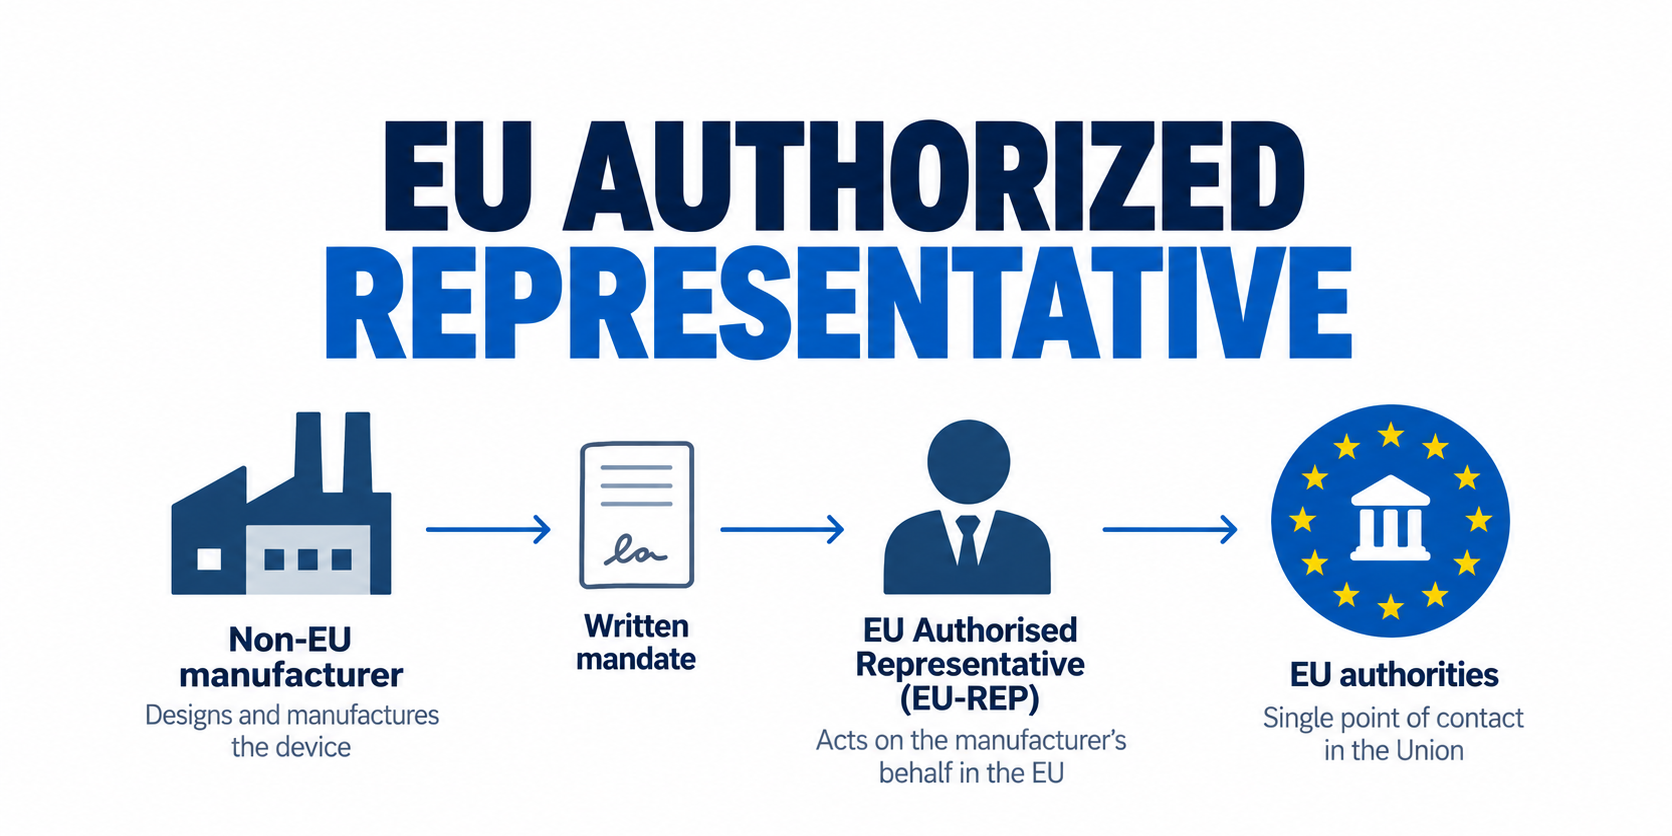
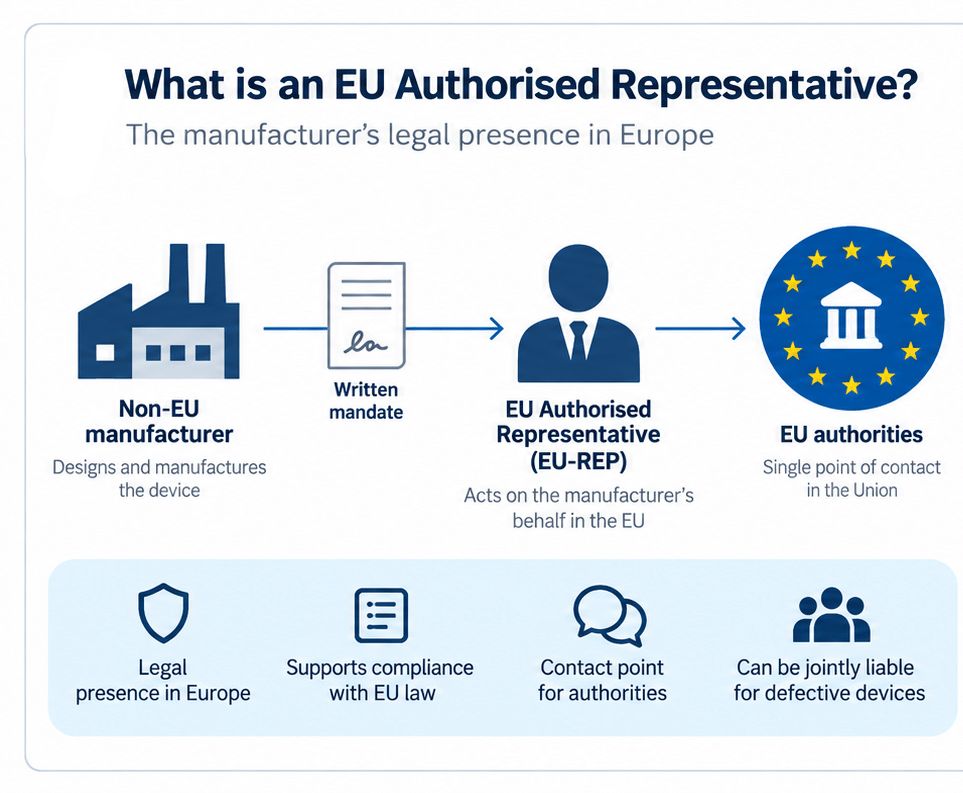
An EU Authorised Representative acts on behalf of a medical device manufacturer based outside the European Union. Established within the EU, the representative serves as a point of contact for the authorities and has specific legal duties under European medical device law.
The requirement is set out in Article 11 of the Regulation (EU) 2017/745 on medical devices, known as the Medical Device Regulation (MDR) and the Regulation (EU) 2017/746 on in vitro diagnostic medical devices, the IVDR. Both require manufacturers established outside the EU to appoint an authorised representative before placing their devices on the EU market. This article focuses on authorised representatives under Article 11 of the MDR and IVDR. Devices intended for clinical investigations or performance studies are subject to separate legal-representation requirements. Specific provisions apply to custom-made devices, so not every documentation or registration requirement described below applies to those devices in the same way.
The relationship between manufacturer and representative is governed by a written mandate. This sets out the representative’s duties and becomes valid once the representative has accepted it in writing. The mandate is required even when both parties belong to the same corporate group, according to the Medical Device Coordination Group’s guidance MDCG 2022-16 on Authorised Representatives. In addition, when registering as an actor in EUDAMED, a non-EU manufacturer must submit a mandate summary document identifying its appointed authorised representative and summarising the mandate. The summary is required for EUDAMED registration and does not replace the full written mandate.
The regulations specify eight minimum tasks that the mandate must cover. These include checking that the required compliance documents exist, keeping copies available for the authorities, responding to regulatory requests and cooperating on safety measures. The manufacturer and representative may also agree on additional services, such as submitting a conformity assessment application to a notified body on the manufacturer’s behalf. The authorised representative must also provide a copy of the mandate to the competent authority upon request.
The mandate must cover at least all devices within the same generic device group. A generic device group is a set of devices having the same or similar intended purposes or a commonality of technology allowing them to be classified in a generic manner without reflecting specific characteristics. A manufacturer can appoint different representatives for different device groups, but must designate only one representative for each group.
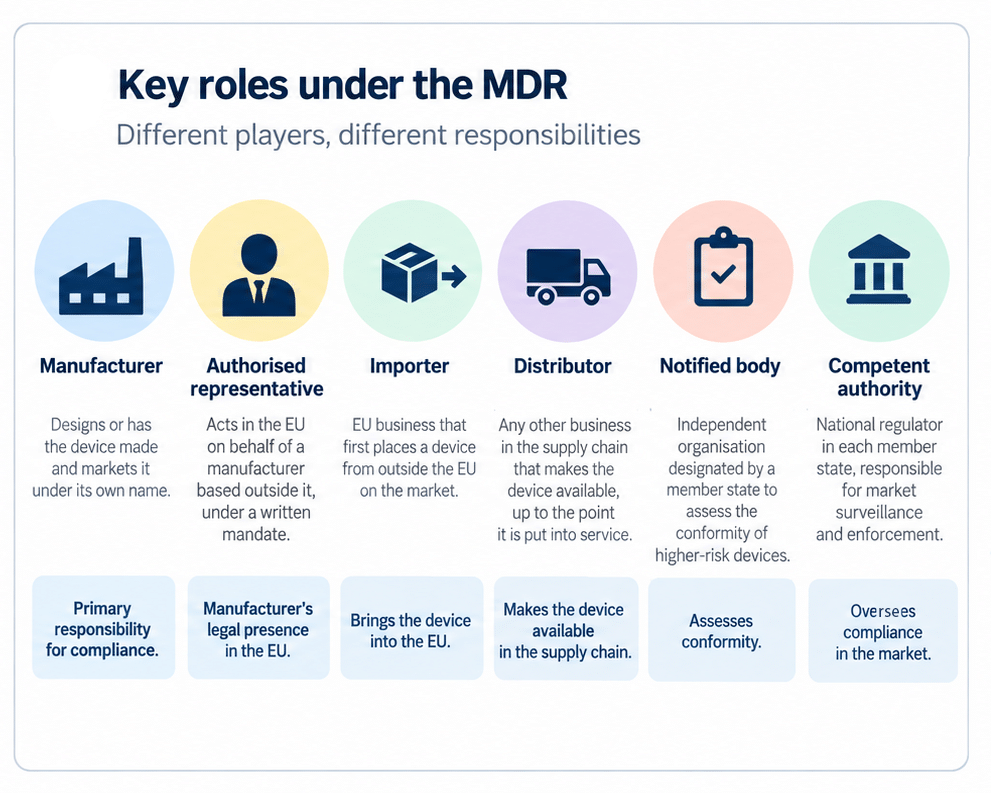
An authorised representative does not replace the manufacturer or take over its core legal obligations. Under Article 11(4) of the MDR and IVDR, the mandate cannot delegate specified manufacturer responsibilities under Article 10. These include obligations relating to design and manufacture, risk management, clinical evaluation under the MDR or performance evaluation under the IVDR, technical documentation, and other specified compliance activities. The representative may assist with these activities, but the manufacturer remains legally responsible for them.
The role is also distinct from that of an importer. An importer is the EU-based business that places a device from outside the EU on the EU market. A company may act as both authorised representative and importer, but each role carries separate legal obligations.
The EU authorised representative role carries specific statutory duties, including the requirement to have a person responsible for regulatory compliance (PRRC) permanently and continuously at its disposal, and potential joint and several liability under the conditions set out in Article 11(5) of the MDR and IVDR.
What responsibilities does an EU-REP have?
The MDR and IVDR each set out eight mandatory tasks that must be included in the written mandate. The manufacturer must provide the EU-REP with the documentation and resources needed to fulfil these responsibilities. The mandate may also cover additional services, such as submitting an application to a notified body on the manufacturer’s behalf.
| Minimum task | What it means in practice | MDR / IVDR |
| Check the paperwork | Confirm that the EU declaration of conformity and technical documentation exist and, where required, that the correct conformity assessment was carried out | Art. 11(3)(a), MDR and IVDR |
| Hold the files | Keep copies of the technical documentation, EU declaration of conformity and, where applicable, relevant certificates, including amendments and supplements, available to competent authorities. Under the MDR, retain these for at least 10 years after the last device covered by the declaration has been placed on the market, or at least 15 years for implantable devices. Under the IVDR, retain these for at least 10 years. | Article 11(3)(b), both regulations; Article 10(8) MDR / Article 10(7) IVDR |
| Register and verify | Comply with its own actor-registration obligations and verify the manufacturer’s compliance with the applicable UDI and device-registration obligations. | Article 11(3)(c), both regulations; Articles 27, 29 and 31 MDR / Articles 24, 26 and 28 IVDR |
| Answer regulators | On request from a competent authority, provide all information and documentation necessary to demonstrate the device’s conformity, in an official Union language determined by the Member State concerned. | Art. 11(3)(d), MDR and IVDR |
| Handle sample requests | Forward to the manufacturer requests for samples or access to a device from the competent authority of the Member State in which the representative has its registered place of business, and verify that the authority receives the samples or is given access. | Art. 11(3)(e), MDR and IVDR |
| Cooperate on safety action | Work with authorities on preventive and corrective measures | Art. 11(3)(f), MDR and IVDR |
| Relay complaints | Tell the manufacturer immediately about complaints and suspected incident reports from clinicians, patients and users | Art. 11(3)(g), MDR and IVDR |
| Terminate the mandate where required | End the mandate if the manufacturer breaches its obligations | Art. 11(3)(h), MDR and IVDR |
Beyond the tasks set out in the mandate, the representative is also in the authorities’ line of sight when they carry out market surveillance. Regulators may consult it about devices that present a serious risk or other non-compliance, require it to hand over documents and information, and inspect its premises, with or without notice.
EUDAMED and changing an EU authorised representative
EUDAMED actor registration does not end with the first entry. The authorised representative must update changes to its own actor-registration information within one week. It must confirm the accuracy of that information no later than one year after initial submission and every second year thereafter, under Article 31(4)-(5) MDR and Article 28(4)-(5) IVDR.
Mandatory use of the first four EUDAMED modules began on 28 May 2026: actor registration, UDI/device registration, notified bodies and certificates, and market surveillance. Applicable transitional arrangements must also be considered.
The representative’s name and address must appear on the device label and in the EU declaration of conformity. A change of representative therefore requires the relevant information to be updated. Under Article 12 of the MDR and IVDR, the arrangements must be defined in an agreement between the manufacturer, the incoming representative and, where practicable, the outgoing representative. The agreement must address the termination and commencement dates, the date until which the outgoing representative may appear in information supplied by the manufacturer, the transfer of documents, and the forwarding of complaints or reports of suspected incidents received after the change.
If the representative ends the mandate because of a breach, it must immediately tell its national authority and, where relevant, the notified body, and give its reasons. As explained in MDCG 2022-16, termination generally applies to the entire generic device group rather than selected devices within it, subject to the qualification concerning devices removed or withdrawn from the market.
Liability
Under Article 11(5) of the MDR and IVDR, where a manufacturer established outside the EU has failed to comply with its Article 10 obligations, the authorised representative is legally liable for defective devices on the same basis as, and jointly and severally with, the manufacturer.
MDCG 2022-16 explains that this liability is conditional on the manufacturer’s liability for the defective device being established under applicable Union or national law, and on the manufacturer’s failure to comply with Article 10. Questions concerning liability are ultimately for the competent courts to determine. MDCG guidance is not legally binding.
The representative therefore has a particular interest in verifying that the manufacturer has fulfilled its obligations, including the requirement to have measures in place to provide sufficient financial coverage for potential liability under Article 10(16) MDR or Article 10(15) IVDR. Such measures may include appropriate liability insurance.
What qualifications does an EU Authorised Representative need to have?
An authorised representative must be established in the EU and comply with the applicable EUDAMED actor-registration requirements. Under Article 15(6) of the MDR and IVDR, it must have a person responsible for regulatory compliance (PRRC) permanently and continuously at its disposal. The PRRC must possess the requisite expertise regarding the regulatory requirements for medical devices or IVDs, as applicable, and satisfy one of the following qualification routes.
- A diploma, certificate or other evidence of formal qualification awarded on completion of a university degree, or a course of study recognised as equivalent by the relevant Member State, in law, medicine, pharmacy, engineering or another relevant scientific discipline, plus at least one year of professional experience in regulatory affairs or quality management systems relating to medical devices under the MDR or IVDs under the IVDR; or
- At least four years of professional experience in regulatory affairs or quality management systems relating to medical devices under the MDR or IVDs under the IVDR.
The PRRC does not have to be an employee, but contractual arrangements must ensure that the person is permanently and continuously at the representative’s disposal. MDCG guidance states that the authorised representative’s PRRC and the non-EU manufacturer’s PRRC should not be the same individual.
A PRRC is not required for representation activities relating solely to legacy devices lawfully placed on the market under the applicable transitional provisions. This does not remove the PRRC requirement for other devices represented under the MDR or IVDR. Applicable legacy-device obligations concerning registration, complaints, vigilance cooperation and market surveillance still apply, and the mandate should reflect those requirements.
How the appointment of an EU-REP works in practice
The following steps outline the appointment process and associated regulatory requirements:
- Sign the mandate: The manufacturer and representative agree a written mandate covering at least the tasks set out in Article 11(3) of the MDR or IVDR, as applicable. The mandate is valid only once the representative has accepted it in writing and must cover at least all devices of the same generic device group under Article 11(2).
- Share the documentation: The manufacturer must ensure that the representative has the necessary documentation permanently available in its current version, under Article 10(8) MDR or Article 10(7) IVDR.
- Register in EUDAMED: The representative must comply with the applicable actor-registration requirements before the devices are placed on the market. Where conformity assessment requires a notified body, the required registration information must be submitted before the application is made. The competent authority validates the registration data and obtains a single registration number (SRN) from EUDAMED for allocation to the representative, under Article 31(1)–(2) MDR or Article 28(1)–(2) IVDR.
- Update the EU declaration of conformity: The declaration must identify the authorised representative by name and registered place-of-business and include its SRN if already issued, under Annex IV, point 1, of the MDR or IVDR.
- Update the device labelling: The representative’s name and registered place-of-business address must appear on the label, under Annex I, Section 23.2(d) MDR or Section 20.2(d) IVDR.
- Verify readiness for placing on the market: Before placing a device on the market, importers must verify that the manufacturer has designated an authorised representative, under Article 13(2)(b) of the MDR or IVDR. Appointment of a representative does not replace the other applicable requirements for placing the device on the market.
- Keep the arrangements current: Changes to the representative’s actor-registration information must be updated within one week, under Article 31(4) MDR or Article 28(4) IVDR. A change of representative requires an agreement addressing the arrangements specified in Article 12 of the applicable regulation.
Elevate Medtech’s EU Authorised Representative services
For manufacturers outside the EU, the right representative can help support a well-managed route to the European market. Elevate Medtech provides EU Authorised Representative services through ELEVATEMEDTECHEU OÜ. This entity accepts the written mandate and performs the authorised representative responsibilities under the applicable MDR or IVDR requirements.
Get in touch with Elevate Medtech to discuss your needs and find out about EU Authorised Representative costs and how we can help.